Medical Devices, Materials, Design & Development, Manufacturing, Quality & Testing, Regulation & Standards
Technical and Regulatory Challenges in Additive Manufacturing of Medical Devices
Updated
Additive manufacturing (AM), often called 3D printing, is steadily moving from prototyping to routine production of medical devices. This transition brings major opportunities, but also introduces new challenges at almost every stage—from design software to final inspection. Engineers, manufacturers, and designers are finding that what worked in traditional device manufacturing does not neatly apply to AM, and that scaling up brings new technical, regulatory, and operational problems. This article highlights practical hurdles specific to additive manufacturing of medical devices and offers clear considerations for overcoming them.
Key Additive Manufacturing Challenges for Medical Devices
AM is not just another way to make existing device designs. Its ability to produce complex, patient-specific components is transforming the field, but also requires manufacturers to rethink workflows, documentation, and quality assurance. The main categories of challenge are:
- Ensuring consistent product quality
- Achieving reproducibility between parts and batches
- Material selection, validation, and reuse
- Workforce skill and design tool limitations
- Aligning with regulatory requirements and standards
- Post-processing, cleaning, and sterilization
Each of these areas needs practical controls and collaboration between engineering, QA, and regulatory teams.
Quality Control and Part Reproducibility
AM processes introduce a high number of variables not present, or much easier to control, in traditional manufacturing. Common quality problems reported include:
- Variability in part dimensions and mechanical properties
- Changes in microstructure due to slight shifts in process parameters
- Batch-to-batch inconsistencies from differences in raw material lots
Ensuring repeatability is difficult because factors such as powder properties, printer calibration, part orientation, and even build location can affect quality. Standard metrology methods may need to be adapted to complex geometries produced by AM. Often, manufacturers must develop a set of critical quality attributes and monitor them as part of the process validation and release.
Process Validation
Regulators (such as the FDA) expect process validation for each AM method used, not just final product testing. This means:
- Defining and locking key process parameters (temperature, build speed, layer thickness, beam power, etc.)
- Documenting the acceptable ranges for these parameters
- Demonstrating—via real data—that chosen parameter ranges reliably produce devices that meet predetermined acceptance criteria
- Validating each step that could affect the final device (e.g., build orientation, removal of support structures, cleaning, post-curing)
Table: Simplified Additive Manufacturing Quality Controls
| Step | Common Quality Risks | Control Strategies |
|---|---|---|
| Pre-print (materials) | Powder/resin purity, degradation | Incoming QC, batch traceability |
| Printing | Parameter drift, equipment issues | Machine qualification, run logs |
| Post-processing | Surface finish, contamination | Validated cleaning, inspection |
| Final inspection | Dimensional errors, mechanical | Metrology, destructive testing |
Material Selection and Validation
Material selection in AM is more complex than in traditional manufacturing. Devices may be made from polymers, metals, ceramics, or even composite materials. Key considerations for medical devices include:
- Biocompatibility (must meet ISO standards or corresponding FDA regulations)
- Stability of properties after exposure to heat, UV, or humidity
- The effect of recycling or reusing powder or resin between builds
AM-specific material risks include:
- Oxidation or moisture absorption during powder storage
- Changes in flow or particle size if reused material is combined with virgin feedstock
Regulators expect that any reuse strategy is validated, with clear evidence that device safety and performance remain within specifications. Manufacturers often need to perform worst-case scenario testing (e.g., maximum-allowed recycled content).
Software, Design, and Digital Workflow Validation
Many AM devices are made starting from a patient’s digital imaging data (CT, MRI). Software used for:
- Segmentation (converting scans to printable files)
- Design (CAD, topology optimization, support structure generation)
- Slicing (turning 3D files into printer instructions)
must all be validated. Any error in these steps can translate into incorrect device geometry or worse, an unsafe product. To ensure reliability, manufacturers should:
- Validate all digital workflows—demonstrate with controlled test cases that the end-to-end process produces accurate, safe device models
- Use version control and electronic recordkeeping to support compliance audits
Post-Processing and Sterilization Barriers
Most 3D printed medical parts require substantial post-processing. Common steps include:
- Removal of support material and excess powder/resin
- Additional machining to meet critical tolerances
- Polishing or smoothing surfaces that will contact tissue
- Thermal or UV curing
- Sterilization
Post-processing steps can introduce new risks:
- Residues or contamination if powders and resins are not fully cleaned
- Loss of dimensional accuracy or undesirable changes to surface properties
- Difficulty sterilizing parts with intricate internal structures or porous surfaces
Post-processing and cleaning must have validated, repeatable procedures with documented acceptance criteria.
Workforce and Design Tool Gaps
One technical issue raised by both industry and regulators is the skills gap. Many design and process engineers are trained for traditional manufacturing but not for AM.
- Understanding how design rules affect printability, surface finish, and part strength is not always intuitive
- Limited familiarity with AM-specific design software
- Few established best practices for designing medical devices for additive processes
Manufacturers report needing to invest in upskilling, recruit AM specialists, or partner with external providers for critical stages.
Regulatory Standards and Alignment Issues
Patchwork of Standards
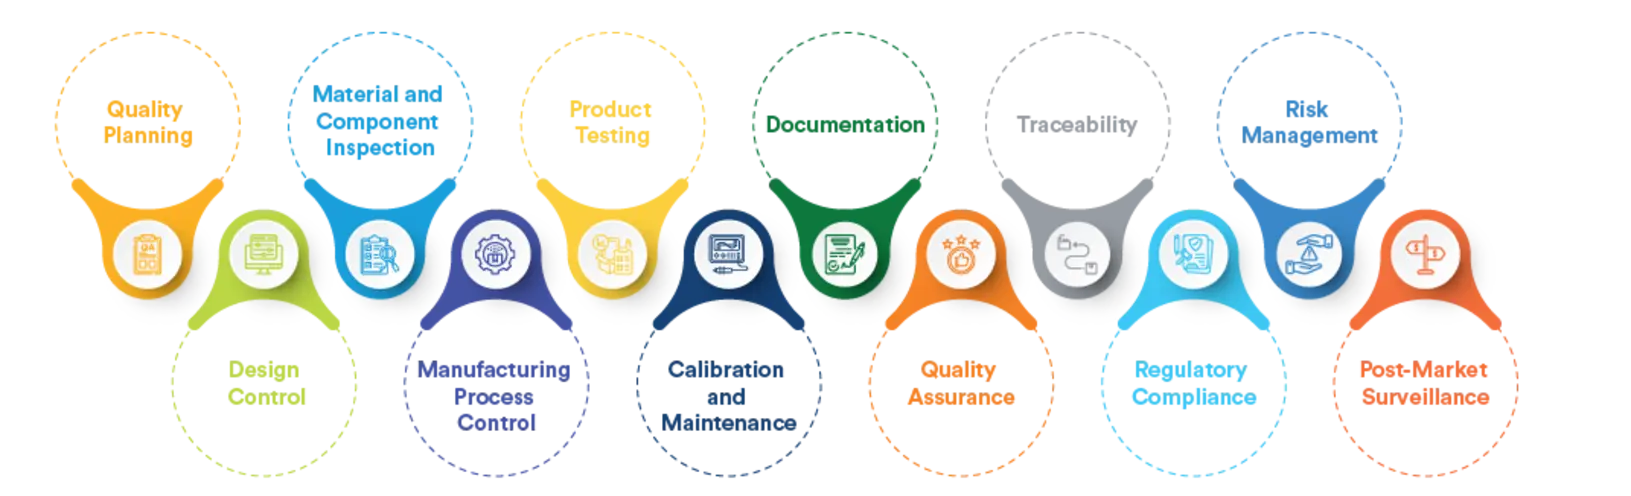
The regulatory environment for AM medical devices is in flux. While core device regulations still apply (such as FDA 21 CFR 820 for Quality System Regulation, and ISO 13485 for quality systems), application of these requirements to additive processes is still evolving.
- Not all international standards (e.g., ASTM, ISO) align fully across countries or with FDA-specific expectations
- Many recognized test standards were written with subtractive or molding processes in mind; they may need adaptation for AM
When a device is custom or patient-specific, premarket pathways can be less clear, especially when clinical data is hard to generalize. Regulators look for:
- Clear evidence of process and product validation
- Quality assurance across the entire digital and physical workflow
- Traceability from raw material to finished device
Point-of-Care Manufacturing
With the rise of in-hospital and point-of-care AM, regulatory questions intensify:
- Who is the “manufacturer”—the hospital, the printer vendor, the device designer?
- How are relevant regulatory responsibilities split and enforced?
Ongoing dialogue between industry, regulators, and academia is needed to clarify and align these requirements.
Manufacturing Scalability and Economics
AM scales well for prototyping and short production runs but often encounters difficulties with:
- High unit costs compared to molding/casting in large volumes
- Throughput limitations due to print speeds or build chamber size
- Additional labor and inspection requirements post-print
AM is especially practical for patient-specific, complex, or low-volume production—but less so for standard, high-volume commodity devices.
Best Practices for Additive Manufacturing Implementation
1. Early and Rigorous Process Validation
- Document critical process parameters and acceptable ranges
- Print validation runs across expected ranges and build locations 2. Material Control and Reuse Management
- Establish clear limits on recycled powder/resin use
- Perform worst-case scenario testing 3. Robust Digital Workflow Validation
- Create and lock validated software toolchains
- Maintain change control and audit trails 4. Documented Post-Processing Protocols
- Validate every cleaning, machining, and sterilization step
- Integrate surface and contamination inspection 5. Invest in Training and Cross-Disciplinary Talent
- Implement targeted training
- Build multidisciplinary review teams 6. Proactive Regulatory Engagement
- Review FDA and other authority guidance early
- Seek feedback prior to submission, especially for custom or novel devices
Practical Summary
Adopting additive manufacturing for medical devices requires more than swapping out equipment. Manufacturers need to build a robust, traceable, and validated process across the entire lifecycle—starting with digital design, through printing, and ending with finished, sterilized, and inspected devices. Each stage introduces new technical and regulatory hurdles that demand specific investment—whether in metrology, documentation, staff skills, or compliance systems. Early, clear validation, and close attention to regulatory guidance, are essential for successful and scalable AM device manufacturing.